Review and verification of technical documentation in accordance with MDR/IVDR Annex II and III, ensuring completeness, consistency, and readiness for regulatory scrutiny.
We support medical device manufacturers in reviewing and strengthening their technical documentation in accordance with MDR/IVDR Annex II and III. Our approach ensures that your documentation is complete, consistent, and ready for regulatory scrutiny, whether for conformity assessment, market access, or ongoing compliance.
Technical documentation is a central element of regulatory compliance and must clearly demonstrate conformity with General Safety and Performance Requirements (GSPR). We perform a structured review of your documentation, identifying gaps, inconsistencies, and areas requiring clarification, while ensuring alignment across all sections.
Our review includes a detailed assessment of labelling and Instructions for Use (IFU), verifying that all mandatory elements are present and compliant with MDR/IVDR Annex I (Chapter III). This includes checks on device identification, intended purpose, warnings and precautions, symbols, language requirements, and the correct inclusion of authorised representative details. We also ensure consistency between labelling, IFU, and the technical documentation.
In addition, we assess key elements such as risk management documentation, clinical evaluation, and post-market surveillance documentation, ensuring coherence and regulatory alignment across the entire technical file.
Our goal is to provide clear, actionable feedback that supports timely compliance and reduces the risk of delays during conformity assessment or regulatory review.
Within this scope, our services include:
- Review of technical documentation against MDR/IVDR Annex II and III
- Gap analysis and identification of missing or inconsistent elements
- Verification of compliance with GSPR (Annex I)
- Review of labelling requirements, including mandatory information and symbol compliance
- Detailed assessment and revision support for Instructions for Use (IFU)
- Consistency checks across technical documentation, labelling, and IFU
- Review of risk management, clinical evaluation, and PMS documentation
- Preparation support for notified body or regulatory submissions
Strengthen Your Technical Documentation
We review your technical documentation against MDR/IVDR requirements to ensure completeness, consistency, and readiness for regulatory scrutiny.
Authorised Representative (EU)
Acting as EU Authorised Representative, including mandate establishment, verification of technical documentation and Declaration of Conformity, and liaison with EU Competent Authorities
EUDAMED Registrations
Management of Actor registration (SRN), UDI-DI and device registration within EUDAMED, ensuring accurate data submission, module alignment, and ongoing maintenance
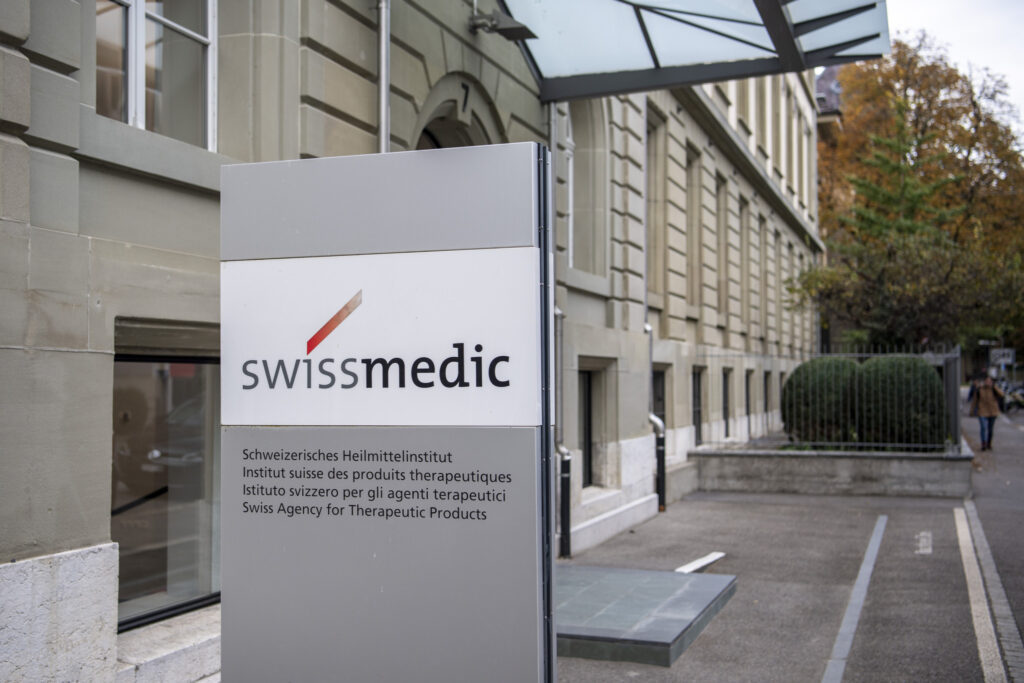
Swiss Authorised Representation
Provision of Swiss Authorised Representative services under Swiss MedDO, including local representation, regulatory verification, and acting as the official contact point for Swissmedic.
Swissmedic Registrations
Preparation and submission of device notifications and economic operator registrations with Swissmedic, ensuring compliance with national regulatory requirements for market access in Switzerland
Regulatory Liaison
Interface with EU Competent Authorities and Swissmedic, including management of regulatory queries, inspections, and formal communications.
Let’s work together
Discover how we support medical device manufacturers across the EU and Switzerland
Trusted Expertise
We work as an extension of your team, providing consistent communication, accountability, and long-term regulatory oversight throughout your product lifecycle.